

Yalantis delivers full-cycle medical device development services for Class II and III products, combining hardware design, firmware engineering, and cloud software under one ISO 13485-certified QMS. From PCB layout to FDA submission, we cut time-to-market by 6 months and reduce post-market costs by 30%.
15+
Years
in healthcare
90%
First-try
approval rate
25+
FDA & CE
approvals
60+
Industry Experts
13485
ISO
Certified QMS
Real projects. Real outcomes.
These projects show what our medical device engineering services look like
in practice, each one with a different device, a different problem, and a different set of regulatory requirements. What stayed the same was the approach: understand the clinical context first, then build something that works in the real world.

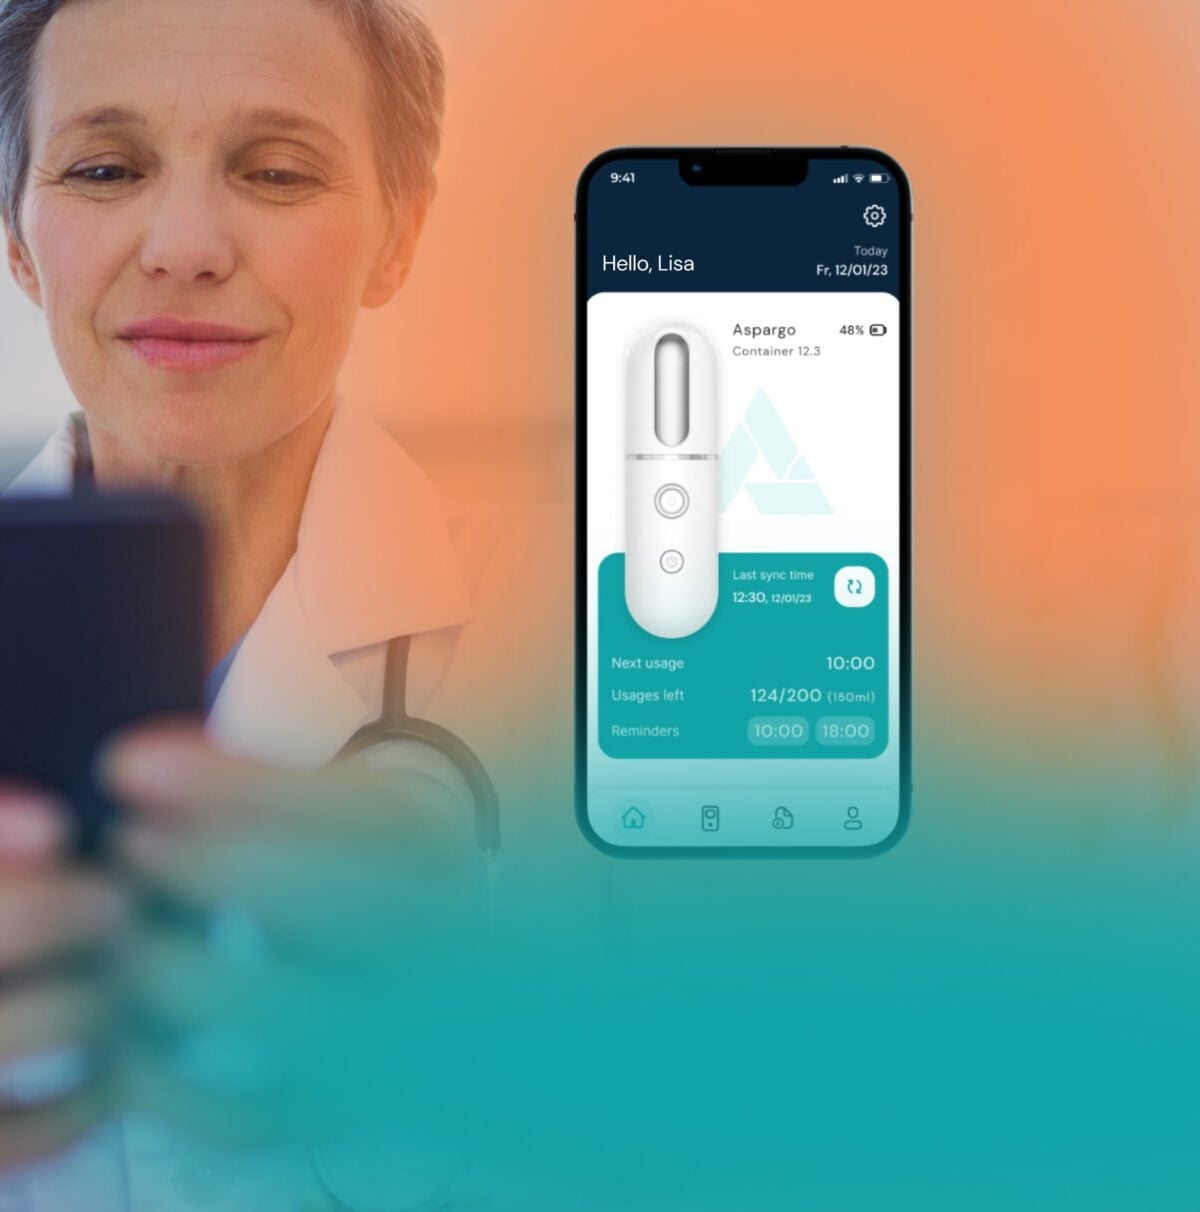
End-to-end medical device engineering services
Most companies split their device work across three or four vendors. We keep it under one roof, from PCB layout to FDA submission, with engineering teams in Europe delivering across the US and EU.
Medical Device design & hardware engineering
- Industrial design and enclosure engineering tailored to clinical use, including human factors engineering
- Consulting on circuit design and electrical architecture
- Rapid prototyping and functional testing of medical equipment
- PCB design compliant with medical-grade requirements
- Component selection with regulatory and certification requirements in mind
- Preparation of full hardware documentation
Medical device firmware development
- Secure embedded application development (Rust, C/C++, MISRA C)
- BSP (Board Support Package) and driver development
- RTOS integration and configuration
- Low-power connectivity implementation (BLE, Wi-Fi)
- Secure bootloader and OTA mechanism setup
Medical device software development
- Mobile applications for device management
- Embedded application development for device control and data acquisition
- SaMD module design and development
Cloud platforms & integrations
- Scalable cloud platforms for real-time data processing
- EHR/EMR integration (HL7, FHIR, DICOM, TEFCA)
- Fleet management and remote provisioning
- HIPAA-compliant backend architecture
AI & TinyML integration
- Development of TinyML models for on-device data processing without internet dependency
- AI algorithm engineering for diagnostics and prediction based on sensor input
- Optimization of machine learning models for constrained hardware
- AI components aligned with FDA guidance on AI-enabled device software functions, including PCCP
- Lifecycle management: continuous learning, validation, and monitoring of AI modules
SaMD development & consulting
- Regulatory strategy and SaMD risk classification (Class I/IIa/IIb/III)
- IEC 62304-compliant architecture and FDA-ready software traceability
- Clinical AI model development with FDA AI/ML compliance
- Algorithm validation, bias mitigation, and FDA Predetermined Change Control Plan (PCCP) alignment
- Patient and clinician app design for diagnostic, monitoring, and therapeutic SaMD, with WCAG 2.2 accessibility compliance
- Cybersecurity-by-design per IEC 81001-5-1
Certification & compliance support
We plan certification from the first sprint, not at the end. All engineering runs under our ISO 13485-certified quality management system, and every hardware, firmware and software deliverable comes with the evidence an FDA reviewer or Notified Body expects. We design, document and test your device against:
- Regulations: FDA 21 CFR 820 (QMSR), FDA 21 CFR Part 11, FDA premarket cybersecurity requirements (Section 524B), EU MDR 2017/745, EU IVDR 2017/746, UK MDR 2002 (UKCA), MDSAP
- Risk, usability and software: ISO 14971, IEC 62366-1, IEC 62304, IEC 82304-1
- Electrical safety, EMC and radio: IEC 60601-1, IEC 60601-1-2, IEC 60529, FCC Part 15, EU RED, RoHS
- Cybersecurity and data protection: IEC 81001-5-1, MDCG 2019-16, HIPAA/HITECH, GDPR
- Interoperability: HL7, FHIR
What we do:
- Regulatory strategy, device classification and certification roadmap (FDA, EU MDR/IVDR, UKCA, MDSAP)
- Design controls per ISO 13485 clause 7.3 and FDA QMSR, with traceability from requirements to risks to tests
- Design and development file (DHF) built and maintained throughout the product lifecycle
- Risk management file (ISO 14971) and usability engineering file (IEC 62366-1)
- Design verification and validation, plus pre-compliance testing for IEC 60601-1 and IEC 60601-1-2 before formal lab testing
- Cybersecurity documentation: threat model, SBOM, security risk assessment, penetration testing
- Technical documentation for 510(k), De Novo, PMA and EU MDR/IVDR; support with FDA and Notified Body questions
- Support for clinical investigations and compliance training
- Post-market: change impact assessments, software updates, regulatory monitoring
- EU Cyber Resilience Act readiness for connected products outside MDR/IVDR scope (wellness wearables, accessories, hospital IoT)
Manufacturing support
- Design for manufacturing and design transfer support to contract manufacturers
- Lifecycle strategy for device components (obsolescence management)
- Coordination of pilot runs and low-volume production
- Troubleshooting production challenges and yield issues
- Ongoing assistance with manufacturing audits and compliance updates
Our technologies
Medical device design and development for Class II and III products
Connected diabetes care
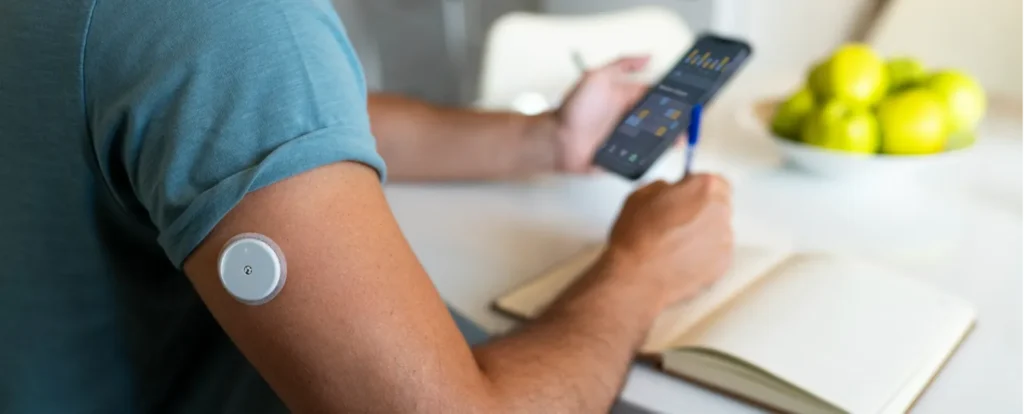
Continuous Glucose Monitors (CGMs), insulin pumps and patch pumps, smart insulin pens, closed-loop artificial pancreas systems, bluetooth-enabled glucometers.
Wearables
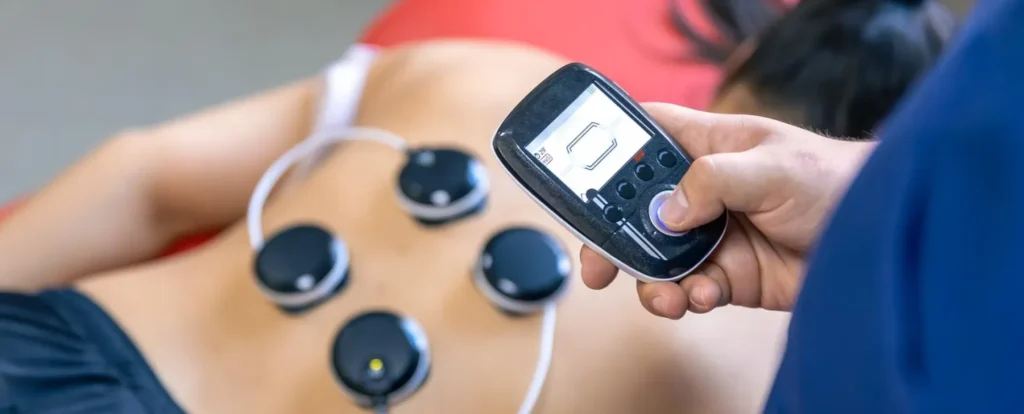
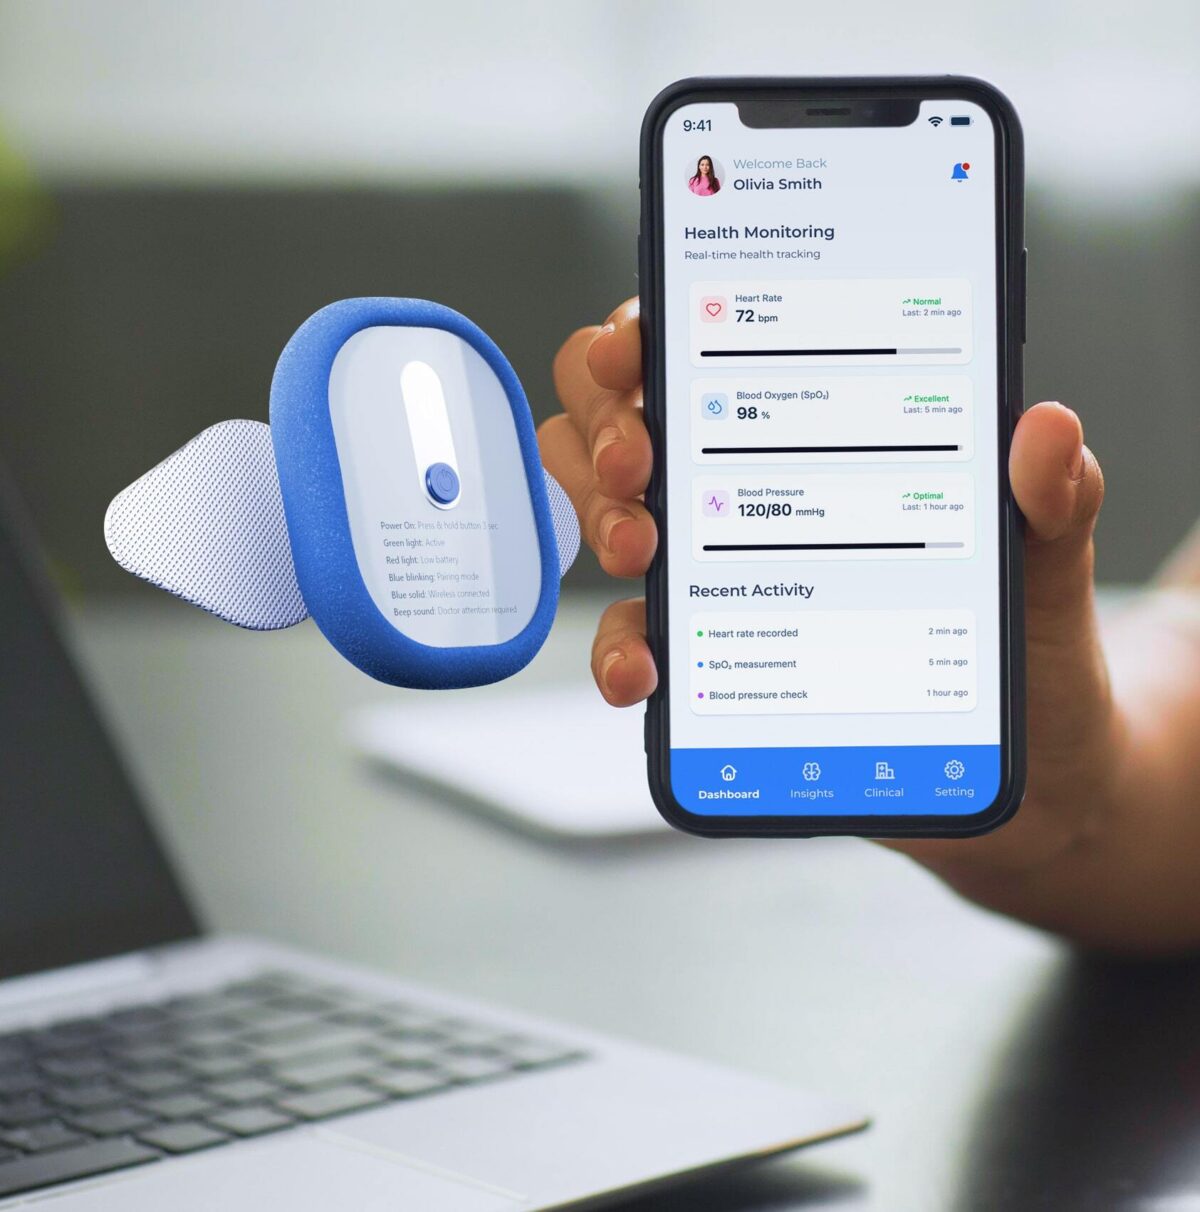
Smart patches for vitals tracking and drug delivery, connected health and fitness trackers, multi-sensor activity monitors for clinical use, ECG-enabled wearables and biosensors.
Cardiac monitoring
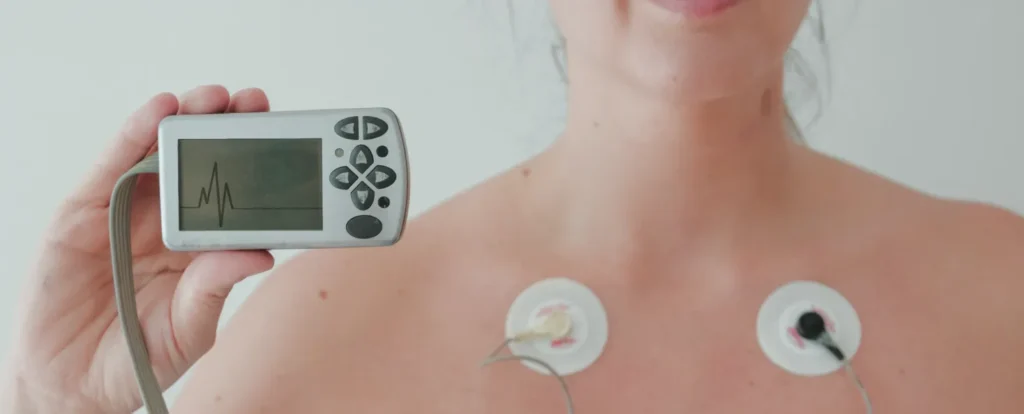
Ambulatory cardiac telemetry devices, implantable cardiac monitors (ICMs), smart holter monitors.
Respiratory devices
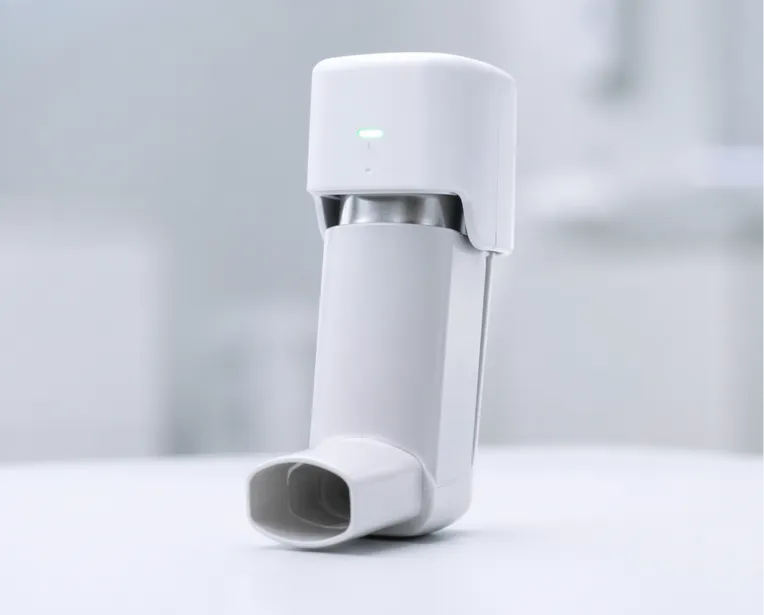
Smart inhalers with adherence tracking, connected CPAP/BiPAP devices, nebulizers with cloud-based data logging, portable spirometers.
Diagnostic equipment
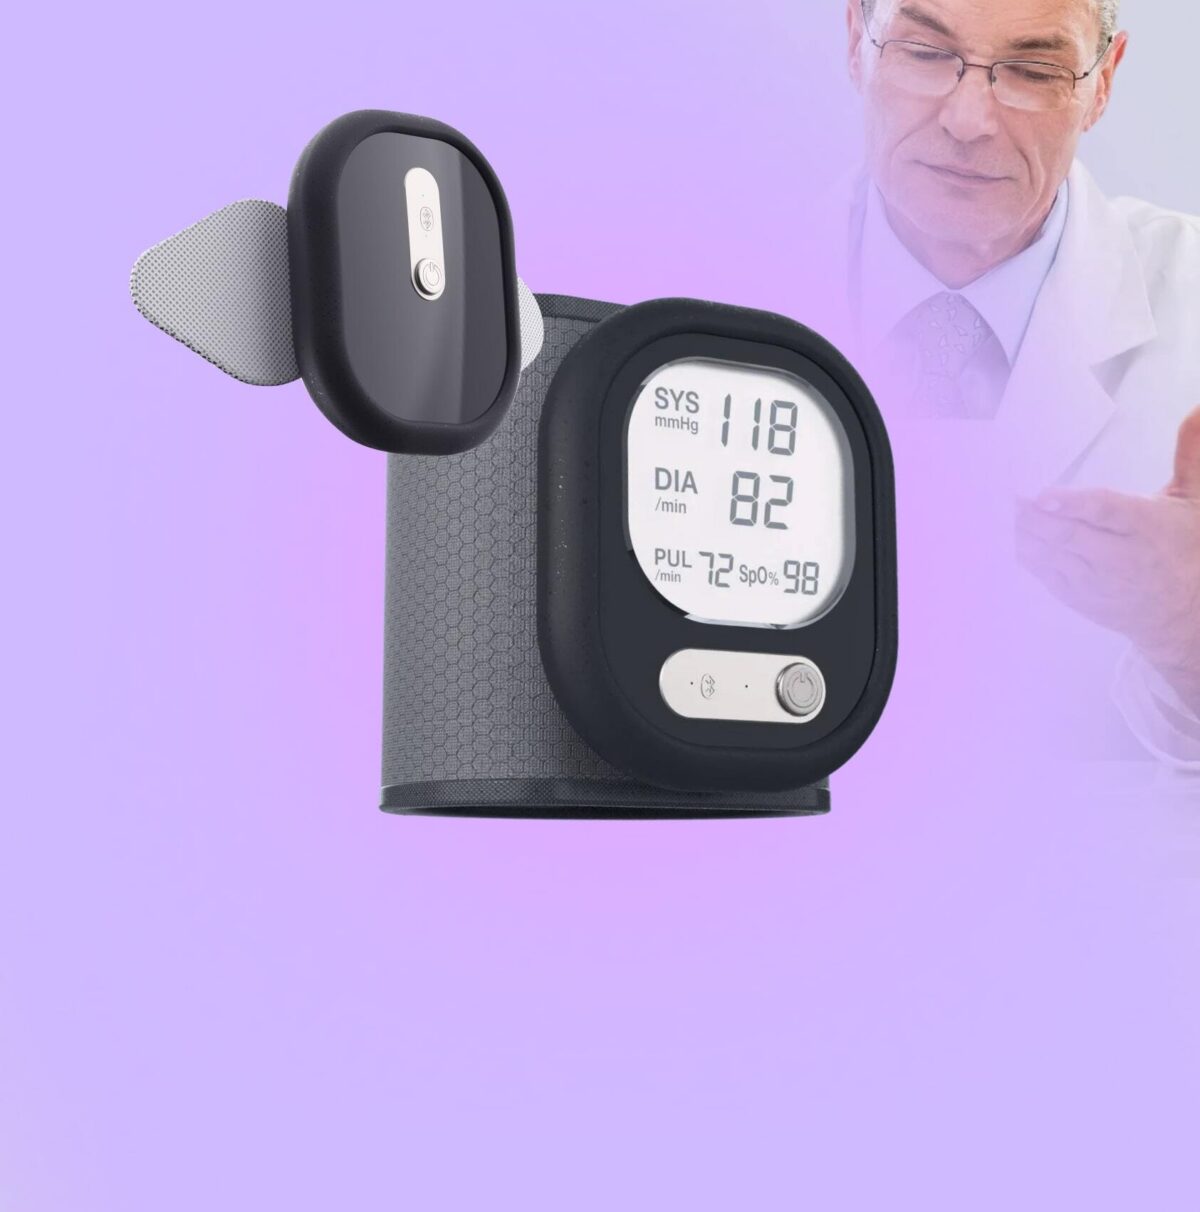
Point-of-care testing (POCT) devices, smart stethoscopes and otoscopes, connected thermometers, oximeters, and vital sign monitors.
Therapeutic & neuromodulation devices
Implantable neurostimulators (spinal cord, vagus nerve), transcutaneous electrical nerve stimulation (TENS) devices, smart orthopedic braces with sensor feedback.
Infusion & drug delivery devices
Smart infusion pumps (syringe, volumetric, elastomeric), implantable infusion systems, electronic auto-injectors, connected wearable injectors.
Hearing & audiology devices
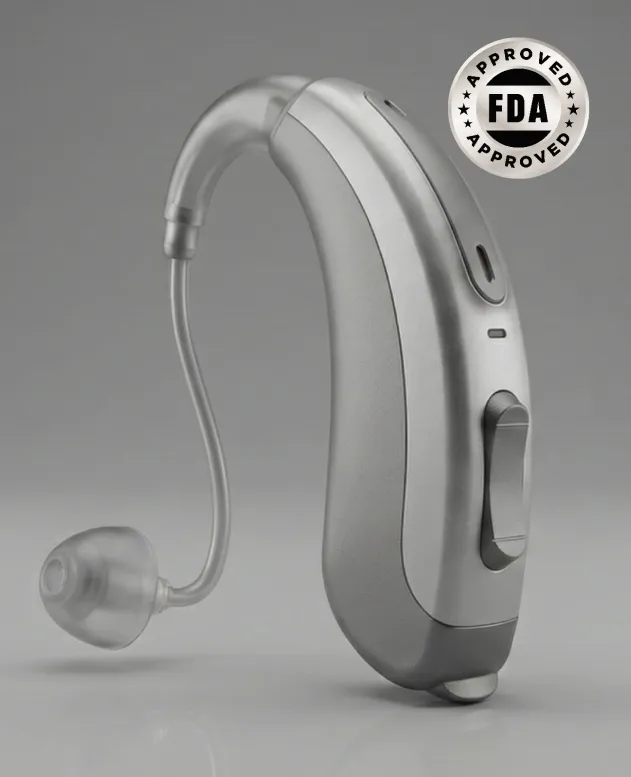
Behind-the-ear (BTE) and OTC hearing aids, app-based hearing aid fitting, connected sound processors, portable audiometers.
Remote patient monitoring hubs & gateways
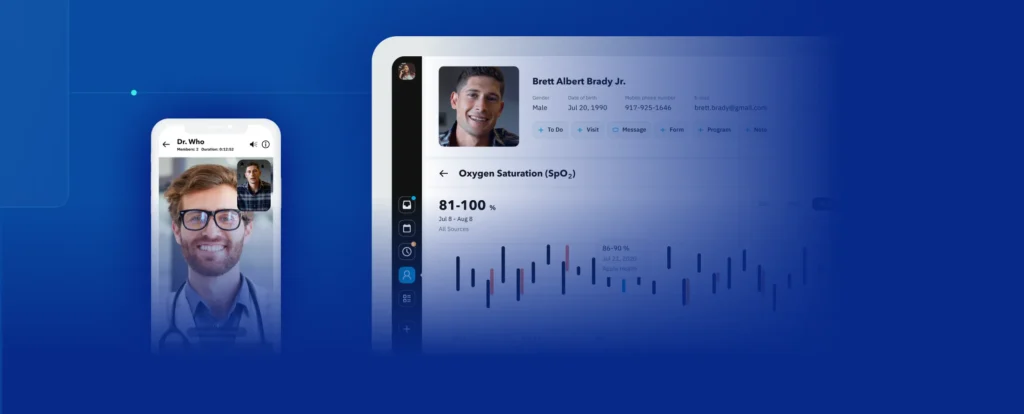
Cellular and Wi-Fi RPM hubs, bedside data gateways, multi-device home monitoring kits, device-to-EHR data bridges.
Medical alert & fall detection
Personal emergency response (PERS) pendants and wristbands, fall detection wearables, panic buttons with location tracking, caregiver alert systems.
Sleep & neurology devices
Home sleep apnea testing (HSAT) devices, EEG headbands and patches, seizure detection wearables, tremor monitoring sensors.
Women’s & maternal health
Wearable fetal and maternal monitors, connected breast pumps, fertility and ovulation trackers, pelvic floor trainers with biofeedback.
Rehabilitation & mobility
Smart orthopedic braces with sensor feedback, sensor-equipped prosthetics, gait analysis insoles, home rehab systems with motion tracking.
Medication management
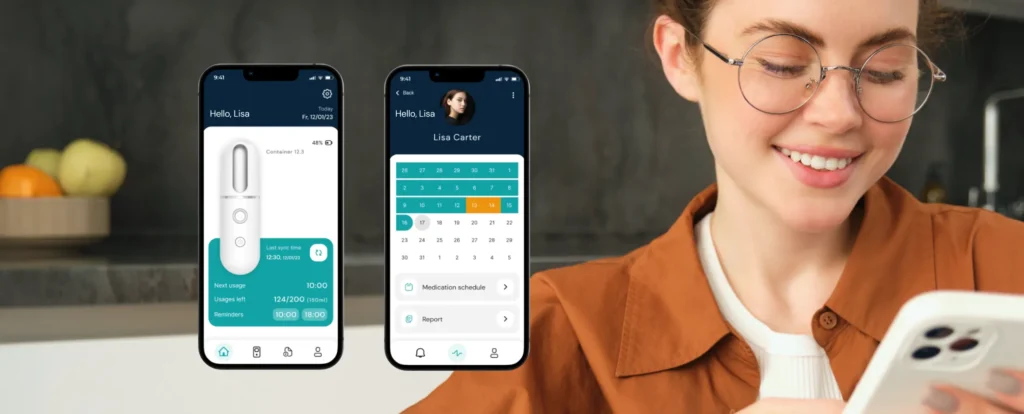
Smart pill dispensers, connected pill bottles and blister packs, add-on sensors for injectors, adherence tracking hubs.
Hospital & clinical equipment
Connected bedside and patient monitors, smart bed sensors, RTLS tags for assets and staff, equipment fleet connectivity.
Want results like these?
Tell us about your project.
Regulatory pathways we support

510(k) premarket notification
The most common route for Class II devices. We prepare your predicate device comparison, performance data, and the full 510(k) submission package so the review goes smoothly.
De Novo classification
For novel low-to-moderate risk devices that lack a valid predicate. We guide you through the De Novo request process, including risk-benefit analysis and proposed special controls.
PMA (Premarket Approval)
Required for Class III, high-risk devices. We support clinical study design, compile the PMA application, and coordinate with the FDA throughout the review cycle.
EU MDR and CE marking
We prepare technical documentation in line with EU MDR 2017/745, coordinate Notified Body interactions, and support CE marking for European market access.
MDSAP
For companies selling into multiple markets, we align your QMS with MDSAP requirements, covering the US, Canada, Brazil, Australia, and Japan in a single audit framework.
Struggling with FDA and ISO requirements?
Get our Guide to ISO 13485 & FDA Compliance for Faster Medical Device Development, and avoid the common mistakes that stall device certification. Includes a detailed roadmap and pro tips.
Our medical device engineering process
Discovery & planning
In the first phase, we run a feasibility assessment and concept development to give you a clear roadmap that reflects your goals and meets regulatory requirements.
Architecture & prototyping
Building on that plan, our engineers deliver early prototypes and system design so you can see how the product will work before full development.
Development in iterations
Once the foundation is set, you’ll see steady progress through iterative design in short cycles, with working features delivered and refined at each step.
Testing & compliance
Alongside the product development process, every stage includes thorough testing and documentation, giving you dependable results and alignment with FDA, MDR, and ISO standards.
Launch & certification
With a fully tested product in hand, our experience with global submissions gets your device to market fully prepared, documented, and supported through approval.
Post-market support
After release and commercialization, you can count on ongoing updates, monitoring, and maintenance to keep your device safe and compliant.
Technologies we work with
Rust
C
C++
Kotlin
Bootloader
Linux Kernel
AWS IoT
Arduino
ESP32
STM32
NRF52
Zephyr
LoRaWAN
MQTT
MISRA C
Yocto
CMSIS
Ready to start?
Schedule a call with our medical device team.
Why partner with Yalantis as your medical device design company
Proven track record
As a medical device engineering company with 15+ years of experience, we deliver medical device design services for Class II and III products. We handle safety-critical complexity where failure is not an option, and we operate as a long-term development partner, not a vendor.
Pioneering Rust expertise for medical devices
Eliminate memory vulnerabilities in device firmware using Rust. We ensure 99.99% uptime and solve safety challenges that generalist dev shops cannot handle.
Rapid medical device engineering staffing
Scale your R&D in 2-4 weeks with specialized medical device engineers. Our team brings deep medical device industry knowledge that stays consistent throughout the device lifecycle.
Audit-ready medical device security & compliance
An ISO 13485-certified partner aligning device architecture with IEC 62304 and FDA standards from day one to streamline your certification.
End-to-end medical device synergy
We bridge hardware and cloud throughout every stage of medical device development, unifying low-level firmware, sensor data, and mobile apps into one seamless connected system.
Engineered for medical device profitability
Maximize margins by optimizing device BOM costs and workflows. We ensure your product makes business sense from prototype to mass production.
Standards and regulations we build your device to
Compliance is part of the engineering work from the first sprint. These are the regulations and standards we design, document and test your device against, so the evidence your FDA reviewer or Notified Body expects grows with the product.
Our certifications
ISO 13485
Medical device quality management,
re-audited annually
ISO 27001
Information security management,
re-audited annually
ISO 9001
Quality management,
company-wide
Standards and certifications we follow
FDA 21 CFR 820 (QMSR)
FDA quality system rule f
or device makers, aligned with ISO 13485.

EU MDR 2017/745
EU regulation for market access
and safety of medical devices.

EU IVDR 2017/746
EU regulation for in-vitro
diagnostic devices, the counterpart to MDR.
UK MDR 2002 / UKCA
UK conformity marking for
medical deviceson the GB market.
ISO 14971
Risk management across
the medical device lifecycle.
IEC 62366-1
Usability engineering applied
to medical devices.
IEC 62304
Software lifecycle for medical device
software and firmware.
IEC 82304-1
Product safety for standalone health
software and companion apps.
FDA 21 CFR Part 11
FDA rule on electronic
records and electronic signatures.
IEC 60601-1
Basic safety and essential performance
of medical electrical equipment.
IEC 60601-1-2
EMC requirements for medical
electrical equipment.
IEC 60529 (IP ratings)
Ingress protection against dust and water.
FCC Part 15
US rules for emissions from
wireless and digital devices.
EU RED 2014/53/EU
EU rules for devices with
Bluetooth, Wi-Fi or cellular radios.
RoHS
EU restriction of hazardous
substances in electronic equipment.
IEC 81001-5-1
Security activities across
the health-software lifecycle.
FDA premarket cybersecurity (Section 524B)
Mandatory FDA cybersecurity evidence
for connected devices, including SBOM.
HIPAA / HITECH
US rules for protecting
Protected Health Information (PHI).
GDPR and UK DPA 2018
EU and UK rules for processing
personal and health data.
HL7 / FHIR
Standards for exchanging
clinical data with EHR systems.
IPC-A-610 (Class 2/3)
Acceptability of electronic
assemblies for high-reliability devices.
IEC 61508
industrial functional safety.
IEC 60601-1
IEC 62304
ISO 14971
IEC 60601-1-2
CE marking under MDR/IVDR
What our clients say
Medical device development insights
Articles
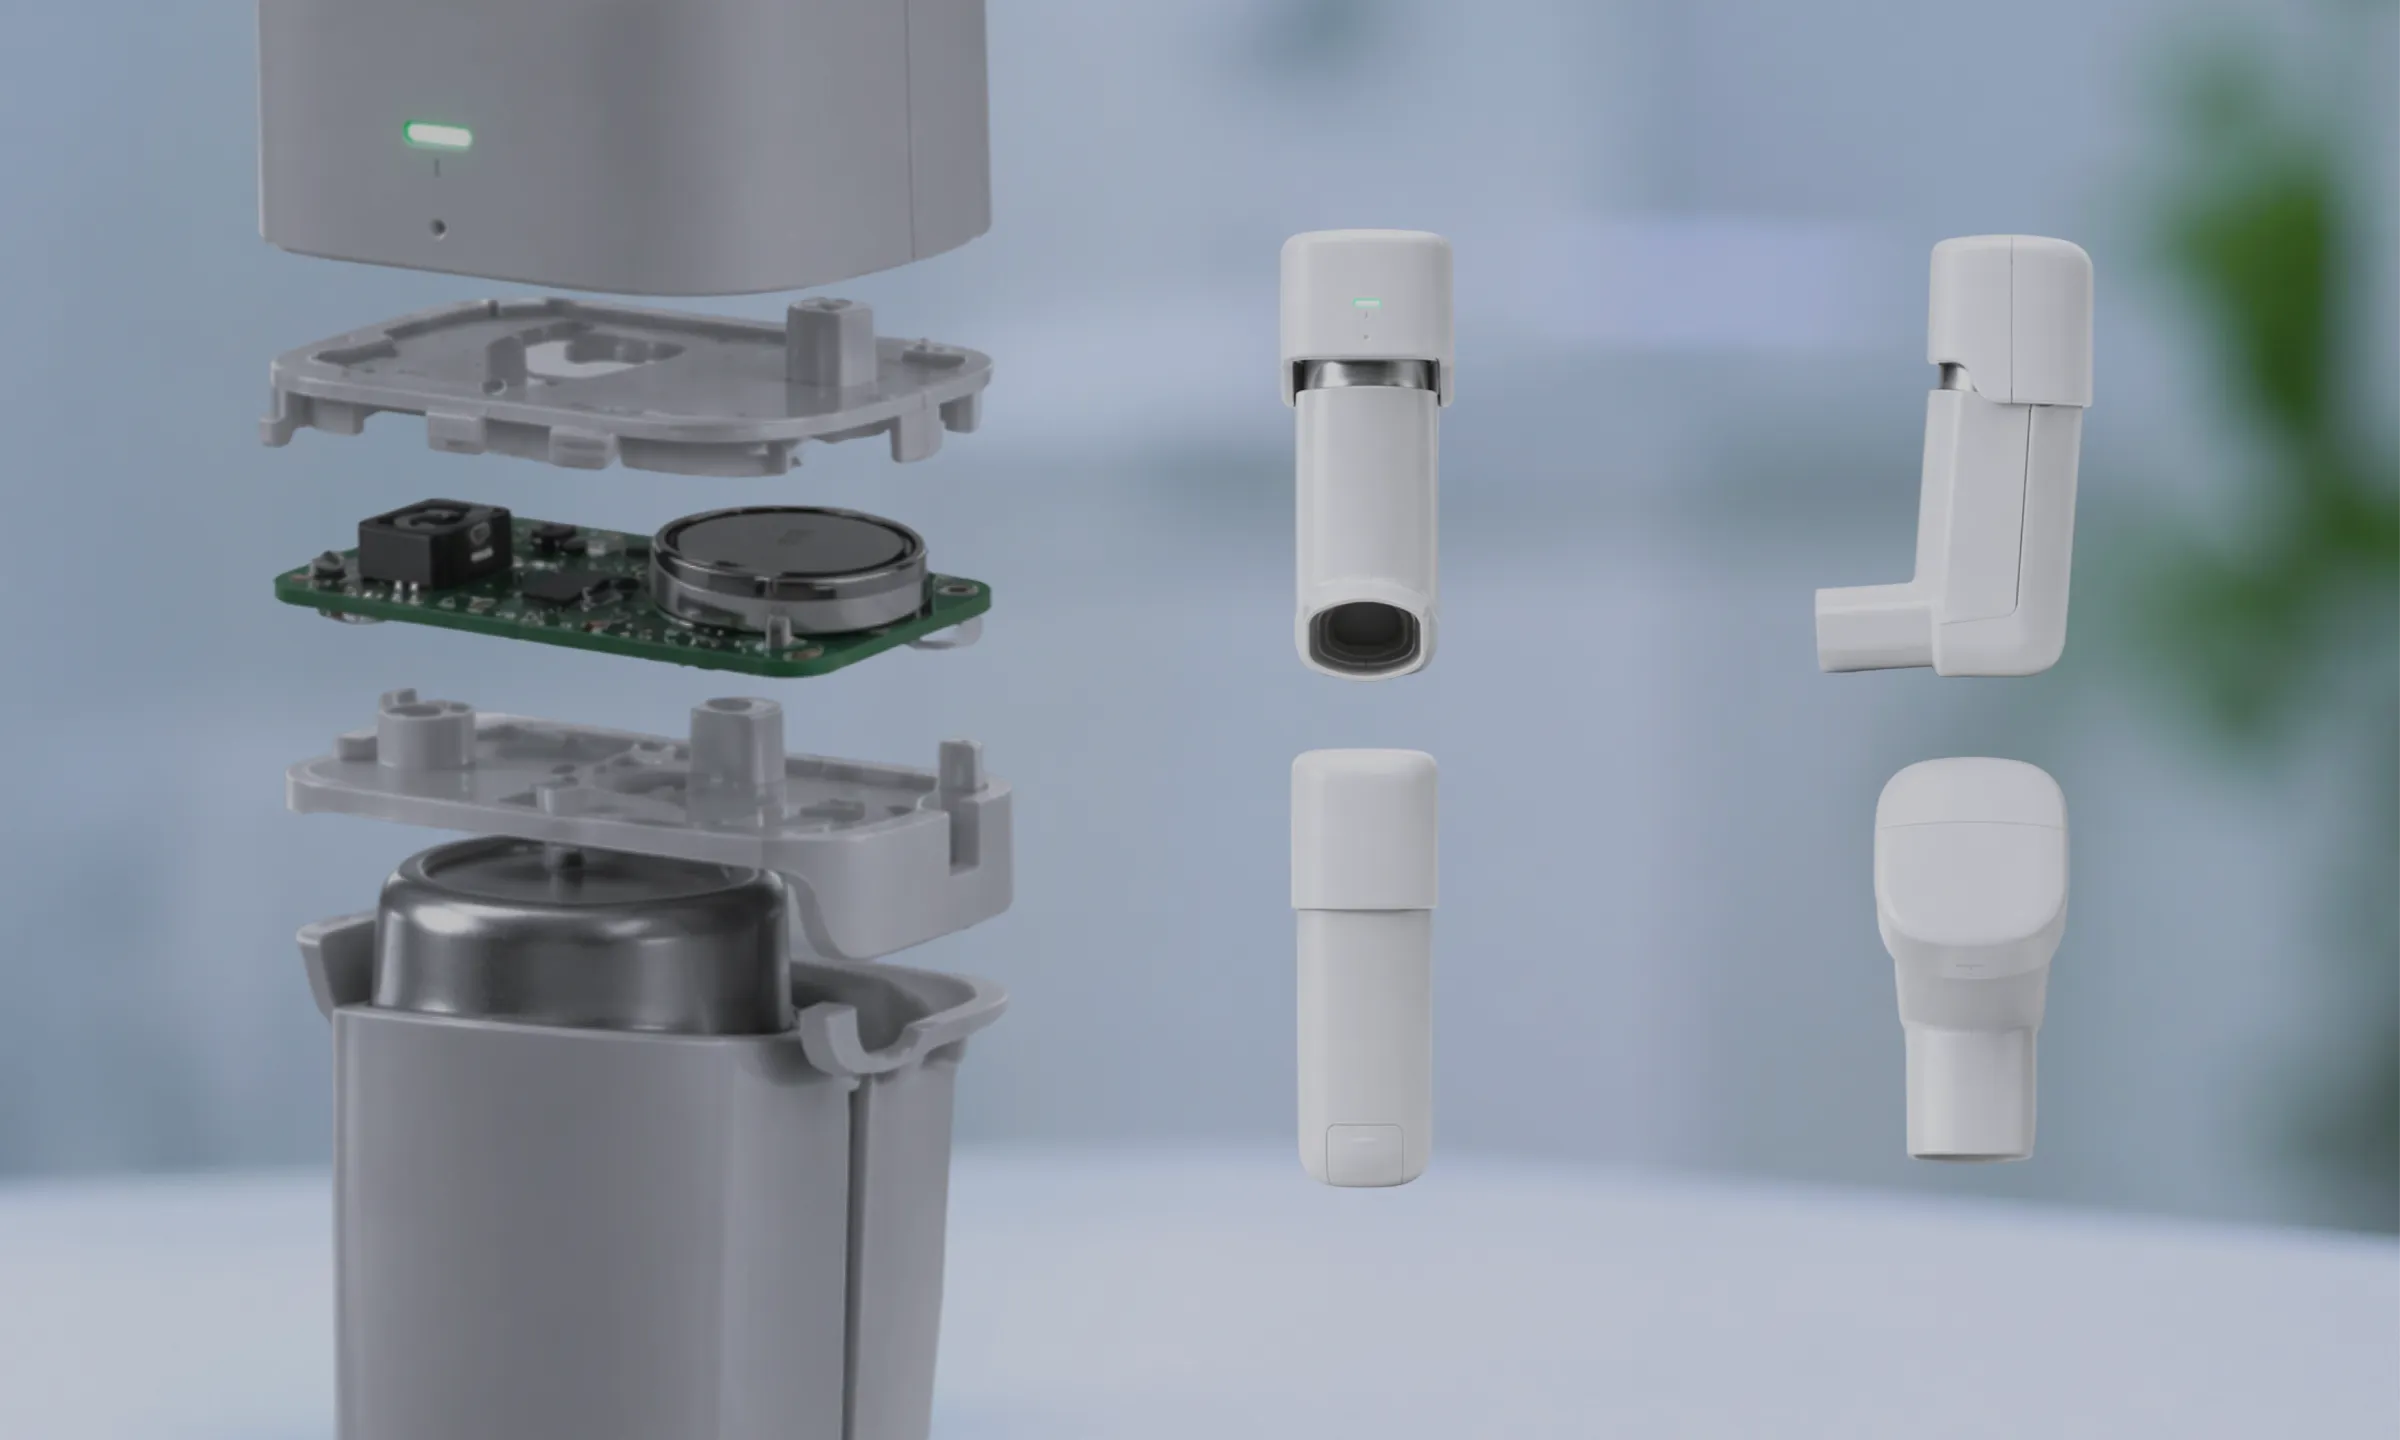
Medical Device Development: The 6 Phases From Concept to Market
Explore the six phases of medical device development, and find out why the first few weeks decide most of your schedule.
Articles
mHealth Strategy in 2026: How to Turn a Patient Engagement App into a SaMD Product
If your patient app doesn’t deliver outcomes, it won’t get paid or adopted. Here’s how to turn it into a regulated, revenue-generating SaMD product.
Articles
A Guide to IEC 62304 & ISO 13485 for Embedded Medical Device Software
If you’re preparing to scale an embedded medical device, this guide breaks down how IEC 62304 and ISO 13485 fit into your workflow and what you need to stay compliant as you grow.
FAQ
-
Do you provide hardware design and PCB engineering, or only software?
We do both. Our medical device engineering services cover the full stack, from PCB design and enclosure engineering to firmware, embedded software, and cloud platforms. This range of services means one team handles hardware and software together, so nothing gets lost in handoffs between vendors.
-
Do you develop Software as a Medical Device (SaMD)?
That’s what we do! We offer medical device software engineering for SaMD products, from risk analysis and UI design to cloud connectivity and compliance.
-
What certifications and standards does Yalantis follow for medical device development?
We operate under an ISO 13485-certified quality management system and are also ISO 9001 and ISO/IEC 27001 certified. On projects, we design and document devices to FDA 21 CFR 820 (QMSR), EU MDR and IVDR, ISO 14971 for risk management, IEC 62304 for software lifecycle, IEC 62366-1 for usability, IEC 60601-1 and IEC 60601-1-2 for electrical safety and EMC, and IEC 81001-5-1 plus FDA premarket cybersecurity requirements for connected devices.
-
Which part of the device does Yalantis build?
We engineer the whole device except serial production: hardware, firmware, software and cloud, plus the documentation that proves they are safe. You can engage us for the full device or for a single layer:
- Hardware: industrial design and enclosure, electronics and PCB design, component selection, prototypes, hardware documentation
- Firmware: embedded software in Rust and C/C++, BSP and drivers, RTOS, BLE/Wi-Fi connectivity, secure boot and OTA updates
- Software: companion mobile apps, SaMD modules, patient and clinician interfaces
- Cloud and integrations: device-to-cloud platforms, fleet management, HL7/FHIR integration with EHRs
- AI: on-device (TinyML) and cloud algorithms, validation, PCCP
- Manufacturing: design for manufacturing, design transfer to your contract manufacturer, pilot runs
We don’t do serial production (your contract manufacturer does), formal certification testing (accredited labs do), and we don’t act as the legal manufacturer of the device.
-
Which documents does Yalantis hand over to the client?
Everything we produce becomes your property and goes into your design history file and technical documentation. A typical handover for a Class II device includes:
- Design and requirements: user needs, system and software requirements, architecture and detailed design, traceability matrix (requirements ↔ risks ↔ tests)
- Hardware: schematics, PCB layout and Gerber files, BOM with approved alternates, mechanical CAD and drawings
- Firmware and software: source code and build instructions, IEC 62304 documentation set (development plan, safety classification, verification records, release notes), SOUP list, SBOM
- Risk and usability: ISO 14971 risk management file, IEC 62366-1 usability engineering file
- Cybersecurity: threat model, security risk assessment, penetration test report, vulnerability management plan
- Verification and validation: test protocols and reports, pre-compliance test results
- Manufacturing: design transfer package, end-of-line test specifications, DFM report
- Regulatory: draft sections for your 510(k), De Novo or PMA submission (eSTAR), or EU MDR/IVDR technical documentation
Documents follow your eQMS templates, or ours if you don’t have them yet.
-
What does Yalantis’ quality management system (QMS) cover?
Our QMS is certified to ISO 13485 for. It governs how we design and develop your device:
- design and development planning, inputs, outputs, reviews, verification, validation, transfer and changes
- risk management (ISO 14971) and software lifecycle processes (IEC 62304)
- document and record control, configuration management
- control of suppliers and subcontractors (components, PCB assembly, test labs)
- engineer training and competence, internal audits, corrective and preventive actions (CAPA)
Our QMS does not replace yours. As the legal manufacturer, you need your own QMS for production, labeling, complaint handling, post-market surveillance and vigilance. We work as your certified design supplier under a quality agreement, either within our QMS or inside your procedures. Project data is protected under our ISO/IEC 27001-certified information security management system.
-
Who is responsible for what: manufacturer, regulatory consultant, lab, or certification body?
As the manufacturer, you carry legal responsibility for the device. Each of the other parties covers one part of the work:
- Manufacturer (you): intended use and claims, final classification, your own QMS, Declaration of Conformity, submissions to the FDA or a Notified Body, registration (FDA listing, EUDAMED), person responsible for regulatory compliance (PRRC, EU), clinical evaluation, labeling, post-market surveillance and vigilance.
- Yalantis: engineering of the device and the evidence you sign and submit: design documentation, risk and usability files, verification and validation, cybersecurity documentation and submission inputs. We answer technical questions and deficiency letters together with you.
- Regulatory consultant: regulatory strategy sign-off, pathway and predicate selection, FDA Pre-Submission meetings, clinical evaluation report, submission compilation, and US Agent or EU Authorized Representative if you’re based outside that market. Our regulatory specialists can cover strategy and documentation or work alongside your consultant. [CONFIRM what’s in-house]
- Accredited test lab: formal testing and reports for electrical safety (IEC 60601-1), EMC (IEC 60601-1-2), radio (FCC, RED), biocompatibility and IP rating, plus CB test reports. We write the test plan, run pre-compliance tests, support the lab and fix findings.
- Certification body or regulator: in the EU, a Notified Body audits your QMS and technical documentation and issues the CE certificate. In the US, the FDA reviews the device and clears or approves it. In the UK, a UK Approved Body handles UKCA. MDSAP audits are run by MDSAP auditing organizations. None of these bodies consult or design.
-
Can you assist with FDA or MDR compliance for medical device software?
Yes. We provide software design for medical devices in line with FDA and MDR regulations, as well as ISO 13485 and IEC 62304 standards, to support smooth approvals and safe releases. That applies to software development for medical devices as well.
-
Can you support EU MDR compliance and CE marking, not just FDA?
Yes. We prepare technical documentation per EU MDR 2017/745, support Notified Body audits, and help with CE marking for European market access. We also handle multi-market strategies through MDSAP, which covers the US, Canada, Brazil, Australia, and Japan.
-
What is the difference between Class II and Class III medical devices?
Class II covers moderate-risk devices like infusion pumps and powered wheelchairs. Class III is for high-risk products such as implantable pacemakers. For Class II, we typically follow the 510(k) pathway with a 12 to 24 month timeline. Class III requires PMA approval with clinical evidence, and usually takes 3 to 7 years. Our design controls and testing scale with the regulatory requirements of each class.
-
How long does it take to develop a Class II or Class III medical device?
For most Class II devices going through the 510(k) pathway, you can expect 12 to 24 months from concept to clearance. Class III devices that need PMA approval typically take 3 to 7 years because of clinical trial requirements and longer FDA review cycles. We work to shorten those timelines by running hardware and software development in parallel and starting documentation early.
-
How much does medical device development cost, and what affects the timeline?
The biggest factors are device class, regulatory pathway, and how much groundwork is already in place. Class II devices with a clear 510(k) predicate generally move faster and cost less, while Class III products requiring PMA take significantly longer and carry higher costs because of clinical trials and extended review cycles. We shorten development timelines by running hardware, firmware, and software in parallel and starting regulatory documentation from the first sprint. Since every project is different, the best way to get a realistic estimate is to talk to our team about your specific device and regulatory situation.
-
How do you handle integration issues with existing healthcare systems?
Challenges in medical device software development often originate from poor vendor documentation or integrating multiple devices. We handle this by working with vendors directly or suggesting better-documented alternatives. For multi-device setups, we merge and normalize data to ensure smooth processing. Every project is unique, so issues may vary.
-
Can you integrate medical devices with EHR or hospital systems?
We can. Yalantis is a medical device design firm that also delivers medical device interoperability services, supporting integration with EHRs, hospital platforms, and HL7/FHIR standards.
-
How do you ensure the interoperability of connected medical devices?
We develop medical device software solutions that standardize and synchronize data across various devices, so the data exchange is smooth and compliant.
-
Do you provide validation and verification services for medical software?
Yes, we do it as part of our medical device software development services. We conduct strict verification and validation (V&V) testing to ensure software safety, reliability, and compliance.
-
What is your experience with healthcare and medtech interoperability?
We’ve delivered medical device engineering services for all classes of FDA-approved devices and built custom medical device solutions from scratch. Our team develops medical device software for diagnostics, remote monitoring, and chronic disease management, and we offer consulting and ongoing outsourcing when needed. This makes us a long-term partner for healthcare companies that require seamless data integration and dependable outcomes.
Explore related services
Let’s set up a call
Leave your info and a few words about the project. We’ll review it and reach out to book a call.
Welcome to Yalantis, please fill out the form and we’ll get back to you.
Thank you for contacting us.
Keep an eye on your inbox. We’ll be in touch shortly
Meanwhile, you can explore our hottest case studies and read
client feedback on Clutch.